Uploaded February 2023 | Updated September 2026, 2 weeks ago
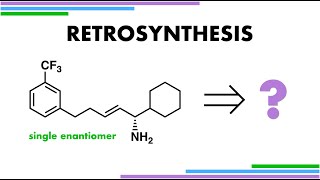
Using asymmetric catalysis for the enantioselective synthesis of this chiral allylic amine, highlighting selectivity issues and rearrangement disconnections.
CBS Reduction Explained:
youtu.be/rv-dZ2qyAzM
More retrosynthesis videos:
youtube.com/playlist?list=PLavaRHHaRimVhyZD79H8g08cfhxrZMcB1
A discussion involving enantioselectivity in hydrogenations in asymmetric catalysis to form chiral amines from enamine intermediates. Why rhodium and ruthenium catalysis with DIPAMP and BINAP ligands, respectively, cannot solve this problem easily.
This retrosynthesis uses a pericyclic sigmatropic rearrangement to simplify the disconnection approach. Specifically, the Overman rearrangement of an allylic alcohol to an allylic amine removes a regioselectivity issue in the synthesis. The Overman rearrangement is a type of [3,3]-sigmatropic rearrangement that can convert a trichloroacetimidate to the corresponding amide with faithful transfer of stereochemistry.
Reframing this retrosynthesis as requiring disconnection of an allylic alcohol makes it much more easy as a problem in organic chemistry as there are many methods for making such a functional group enantioselectively. This video proposed the use of a CBS reduction, a cornerstone of asymmetric catalysis, to generate the chiral allylic alcohol in likely very high enantiomeric excess for the desired configuration at the stereogenic centre (chiral centre).
The precursor for the CBS reductions – an alpha,beta-unsaturated ketone – can be easily synthesised by HWE olefination (Horner-Wadsworth-Emmons reaction), with probably a higher selectivity for the E (trans) geometry of the carbon-carbon double bond.
The starting material requires a simple bromination of a meta-directing substituted benzene ring. Then the aryl bromide product can be made into the Grignard organometallic and converted to the soft cuprate ready for Michael addition (conjugate addition, 1,4-addition) at a soft electrophile.
Using asymmetric catalysis for the enantioselective synthesis of this chiral allylic amine, highlighting selectivity issues and rearrangement disconnections.
CBS Reduction Explained:
youtu.be/rv-dZ2qyAzM
More retrosynthesis videos:
youtube.com/playlist?list=PLavaRHHaRimVhyZD79H8g08cfhxrZMcB1
A discussion involving enantioselectivity in hydrogenations in asymmetric catalysis to form chiral amines from enamine intermediates. Why rhodium and ruthenium catalysis with DIPAMP and BINAP ligands, respectively, cannot solve this problem easily.
This retrosynthesis uses a pericyclic sigmatropic rearrangement to simplify the disconnection approach. Specifically, the Overman rearrangement of an allylic alcohol to an allylic amine removes a regioselectivity issue in the synthesis. The Overman rearrangement is a type of [3,3]-sigmatropic rearrangement that can convert a trichloroacetimidate to the corresponding amide with faithful transfer of stereochemistry.
Reframing this retrosynthesis as requiring disconnection of an allylic alcohol makes it much more easy as a problem in organic chemistry as there are many methods for making such a functional group enantioselectively. This video proposed the use of a CBS reduction, a cornerstone of asymmetric catalysis, to generate the chiral allylic alcohol in likely very high enantiomeric excess for the desired configuration at the stereogenic centre (chiral centre).
The precursor for the CBS reductions – an alpha,beta-unsaturated ketone – can be easily synthesised by HWE olefination (Horner-Wadsworth-Emmons reaction), with probably a higher selectivity for the E (trans) geometry of the carbon-carbon double bond.
The starting material requires a simple bromination of a meta-directing substituted benzene ring. Then the aryl bromide product can be made into the Grignard organometallic and converted to the soft cuprate ready for Michael addition (conjugate addition, 1,4-addition) at a soft electrophile.