Uploaded July 2025 | Updated November 2026, 3 days ago
Speaker: Mariko Okada, Professor at the Institute for Protein Research, Osaka University (Japan).
Patient heterogeneity makes cancer treatment and drug development difficult and costly. Therefore, the development of computational methods to find prognostic markers for individualised treatment and drug screening is urgently needed. To address this problem, we employ both network/pathway-based simulation (mechanistic modelling) and molecular-based simulation (molecular dynamics (MD) simulation).
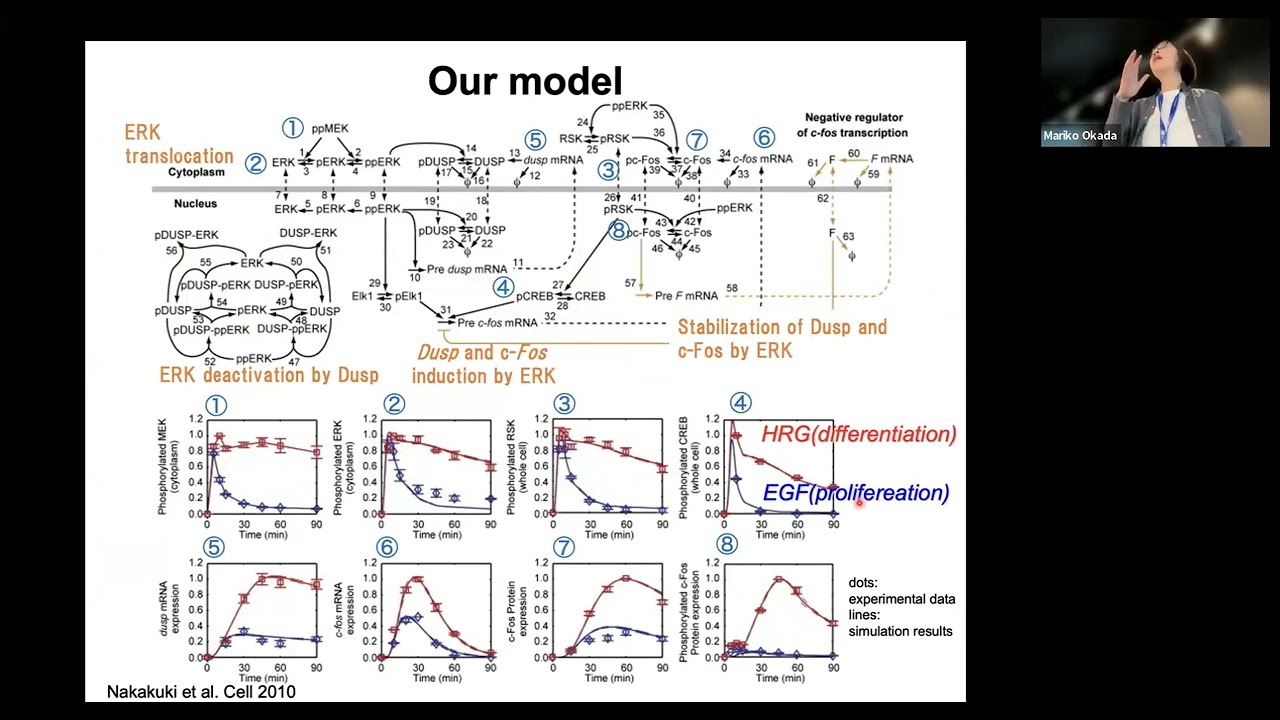
Mechanistic modeling of gene networks using ordinary differential equations (ODEs) is regarded as a promising approach to uncover regulatory mechanisms and identify drug targets in human diseases. We developed Pasmopy (Patient-Specific Modelling in Python), a computational framework for patient stratification based on in silico signalling dynamics. With this framework, we constructed a comprehensive mechanistic model of the ErbB receptor - c-Myc signalling network, trained on phospho-proteomics data from breast cancer cell lines. We then performed a simulation of 377 breast cancer patients using transcriptome data from The Cancer Genome Atlas (TCGA) as the model's initial value. Through this approach, we successfully predicted the key maker genes associated with poor prognosis of triple-negative breast cancer (TNBC) based on the time-course patterns of in silico signalling dynamics, using deep learning method.
In this study, we also developed a new computational tool called Text2model, which transforms the descriptions of biochemical reactions into mechanistic models. Building on Text2Model, we are currently developing natural language processing (NLP) methods to automatically construct mathematical models from the literature and public databases, and to predict gene-drug interactions.
Further information here: bsc.es/research-and-development/research-seminars/sorswomeninbsc-multi-scale-cancer-signaling-network-modeling-using-natural-language-processing
Speaker: Mariko Okada, Professor at the Institute for Protein Research, Osaka University (Japan).
Patient heterogeneity makes cancer treatment and drug development difficult and costly. Therefore, the development of computational methods to find prognostic markers for individualised treatment and drug screening is urgently needed. To address this problem, we employ both network/pathway-based simulation (mechanistic modelling) and molecular-based simulation (molecular dynamics (MD) simulation).
Mechanistic modeling of gene networks using ordinary differential equations (ODEs) is regarded as a promising approach to uncover regulatory mechanisms and identify drug targets in human diseases. We developed Pasmopy (Patient-Specific Modelling in Python), a computational framework for patient stratification based on in silico signalling dynamics. With this framework, we constructed a comprehensive mechanistic model of the ErbB receptor - c-Myc signalling network, trained on phospho-proteomics data from breast cancer cell lines. We then performed a simulation of 377 breast cancer patients using transcriptome data from The Cancer Genome Atlas (TCGA) as the model's initial value. Through this approach, we successfully predicted the key maker genes associated with poor prognosis of triple-negative breast cancer (TNBC) based on the time-course patterns of in silico signalling dynamics, using deep learning method.
In this study, we also developed a new computational tool called Text2model, which transforms the descriptions of biochemical reactions into mechanistic models. Building on Text2Model, we are currently developing natural language processing (NLP) methods to automatically construct mathematical models from the literature and public databases, and to predict gene-drug interactions.
Further information here: bsc.es/research-and-development/research-seminars/sorswomeninbsc-multi-scale-cancer-signaling-network-modeling-using-natural-language-processing